

CE Marking for Electric Toothbrushes: EU Market Entry Guide (2026)
A practical guide to EU CE marking for electric toothbrushes: assess intended purpose, identify applicable requirements, prepare technical documentation and determine whether a notified body is required.
EU Requirements for Electric Toothbrushes: CE Marking and Compliance
To prepare an electric toothbrush for EU sale, identify the product legislation that applies to its design and intended use. Assess conformity, compile the required technical documentation and EU declaration, and affix the CE mark when the applicable route requires it. Keep the model, evidence and market claims within the assessed scope.
An electric toothbrush may be subject to several product rules, depending on its design, power supply, radio functions, materials and intended purpose. The assessment may include electrical safety, electromagnetic compatibility, chemical restrictions and general product-safety obligations. These requirements do not all create separate CE marks, and not every rule applies to every configuration.
Retailers and marketplaces may request product-specific compliance documents as part of their own onboarding or listing policies. Their requirements can differ by product and sales channel; they do not replace the manufacturer's legal assessment.
Use early risk reviews, sample testing and supplier-document checks to identify design issues while the team can still address them. Record findings and corrective actions in the project file.
For oral care brands sourcing from China, agree early with the manufacturer who will provide specifications, component information, samples and other project records. Read our electric toothbrush OEM guide for more on planning a product project.
Which EU Requirements May Apply to Electric Toothbrushes
Start with the product's intended purpose, technical design, power source, radio functions, claims and target markets. Use those facts to identify applicable EU legislation and the conformity-assessment route; do not assume that every electric toothbrush follows the same checklist.
MDR 2017/745 (Medical Device Regulation; when applicable)
Regulation (EU) 2017/745 (MDR) applies when a product meets the legal definition of a medical device, including its manufacturer's stated intended purpose. A toothbrush is not automatically a medical device because it cleans teeth or because marketing mentions plaque. Review the intended purpose, labeling and claims together before deciding whether the MDR applies.
If the MDR applies, classification follows the rules in Annex VIII and the device's intended purpose and characteristics. Some Class I devices may use a manufacturer-led conformity route, while specific Class I categories require notified-body involvement; higher-risk classes generally require a notified body. Do not assign a class from a single marketing phrase alone.
For an MDR device, the manufacturer must meet the requirements for its classification and role, including applicable registration and UDI duties. EUDAMED obligations vary by device and economic operator. Use of the first four EUDAMED modules has been mandatory since 28 May 2026 for the activities covered by those modules.
Electromagnetic Compatibility (EMC)
Assess electromagnetic compatibility for the finished product and its configuration under Directive 2014/30/EU where it applies. The design, charging system, cables and intended operating environment can affect the assessment.
The test plan and standards depend on the product and applicable legislation. A laboratory may evaluate emissions and immunity using standards appropriate to the final configuration and the relevant EU references in force; do not treat one test list as mandatory for every toothbrush.
Assess a charging base or adapter as part of the complete configuration. Radio Equipment Directive 2014/53/EU may apply when the product intentionally uses radio equipment; inductive charging alone does not automatically make a toothbrush a radio product.
Low Voltage Directive 2014/35/EU (within its voltage scope)
Directive 2014/35/EU covers electrical equipment designed for use within 50–1,000 V AC or 75–1,500 V DC. A battery-powered brush may fall below those limits, while a mains adapter or other equipment in the supply chain may need a separate scope assessment.
Assess the charger or adapter against the voltage limits and applicable safety rules. For household oral-hygiene appliances, IEC 60335-2-52 may be relevant alongside IEC 60335-1, subject to the product, applicable edition and EU status of the standard. The test plan should match the equipment actually placed on the market.
A consumer electric toothbrush outside the Low Voltage Directive may still fall under other product-safety legislation. The General Product Safety Regulation (EU) 2023/988 has applied since 13 December 2024 to consumer products in its scope and operates alongside sector-specific rules; it does not itself provide a route to CE marking.
RoHS Directive 2011/65/EU
RoHS Directive 2011/65/EU restricts specified substances in electrical and electronic equipment within its scope. The applicable product category and any exemptions must be assessed; do not assign all toothbrushes to Category 11 without checking the legal classification.
The restricted substances include lead, mercury, cadmium, hexavalent chromium, PBB, PBDE and four phthalates: DEHP, BBP, DBP and DIBP. Limits apply at the homogeneous-material level, subject to the Directive's scope and exemptions. Supplier information, bills of materials and targeted testing can help support the assessment.
For an OEM project, collect current material and component information from relevant suppliers, including for the battery, motor, electronics and housing. Review the evidence against the product's configuration and applicable requirements; a supplier declaration alone may not answer every compliance question.
MDR applicability
Applies only when the product meets the MDR definition of a medical device. Intended purpose and classification determine the conformity route.
EMC requirements
Assess where the Directive applies, based on the finished product and configuration. Select emissions and immunity evidence appropriate to the applicable rules.
Low Voltage requirements
Applies to equipment within the Directive's voltage limits. A mains adapter may need assessment even when the battery-powered brush itself is below those limits.
RoHS scope
Assess whether the equipment falls within RoHS scope, its category, applicable substance limits and any exemptions; document the material-level evidence.
CE Conformity Assessment: A Practical Planning Framework
Set the conformity-assessment sequence from the product’s intended purpose, final configuration, target markets and applicable legislation. Compile technical information while testing plans and evidence are developed.
Identify Applicable Requirements
Document the product configuration, intended purpose, claims, target markets and applicable legislation. Identify the relevant conformity-assessment route and standards, then record the reasoning and any open questions.
Conduct Risk Analysis
Review foreseeable hazards such as battery and charging risks, liquid ingress, brush-head detachment, electrical or mechanical faults, and misuse during cleaning or charging. Link each hazard to a design control or verification step, using a risk-management method appropriate to the product and its regulatory route.
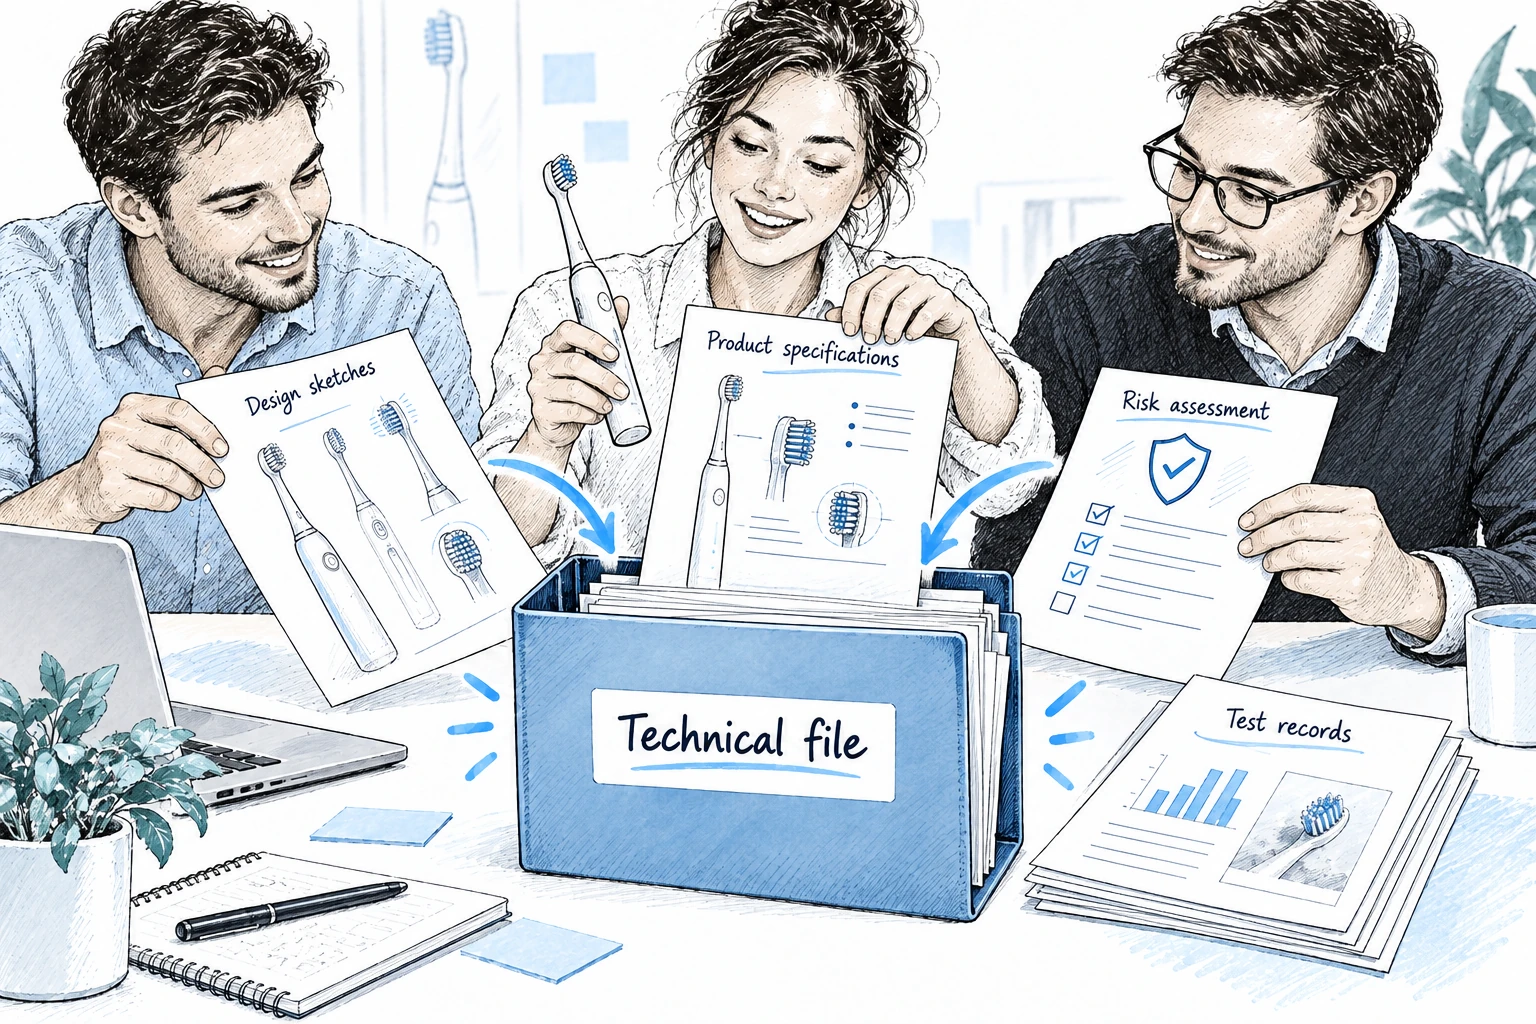
Develop Technical File
Build the technical documentation as the design and evidence develop. Depending on the applicable rules, it may include product specifications, drawings, a bill of materials, risk records, supplier information, test evidence, labeling and the required declarations. Do not wait for every test report before beginning the file.
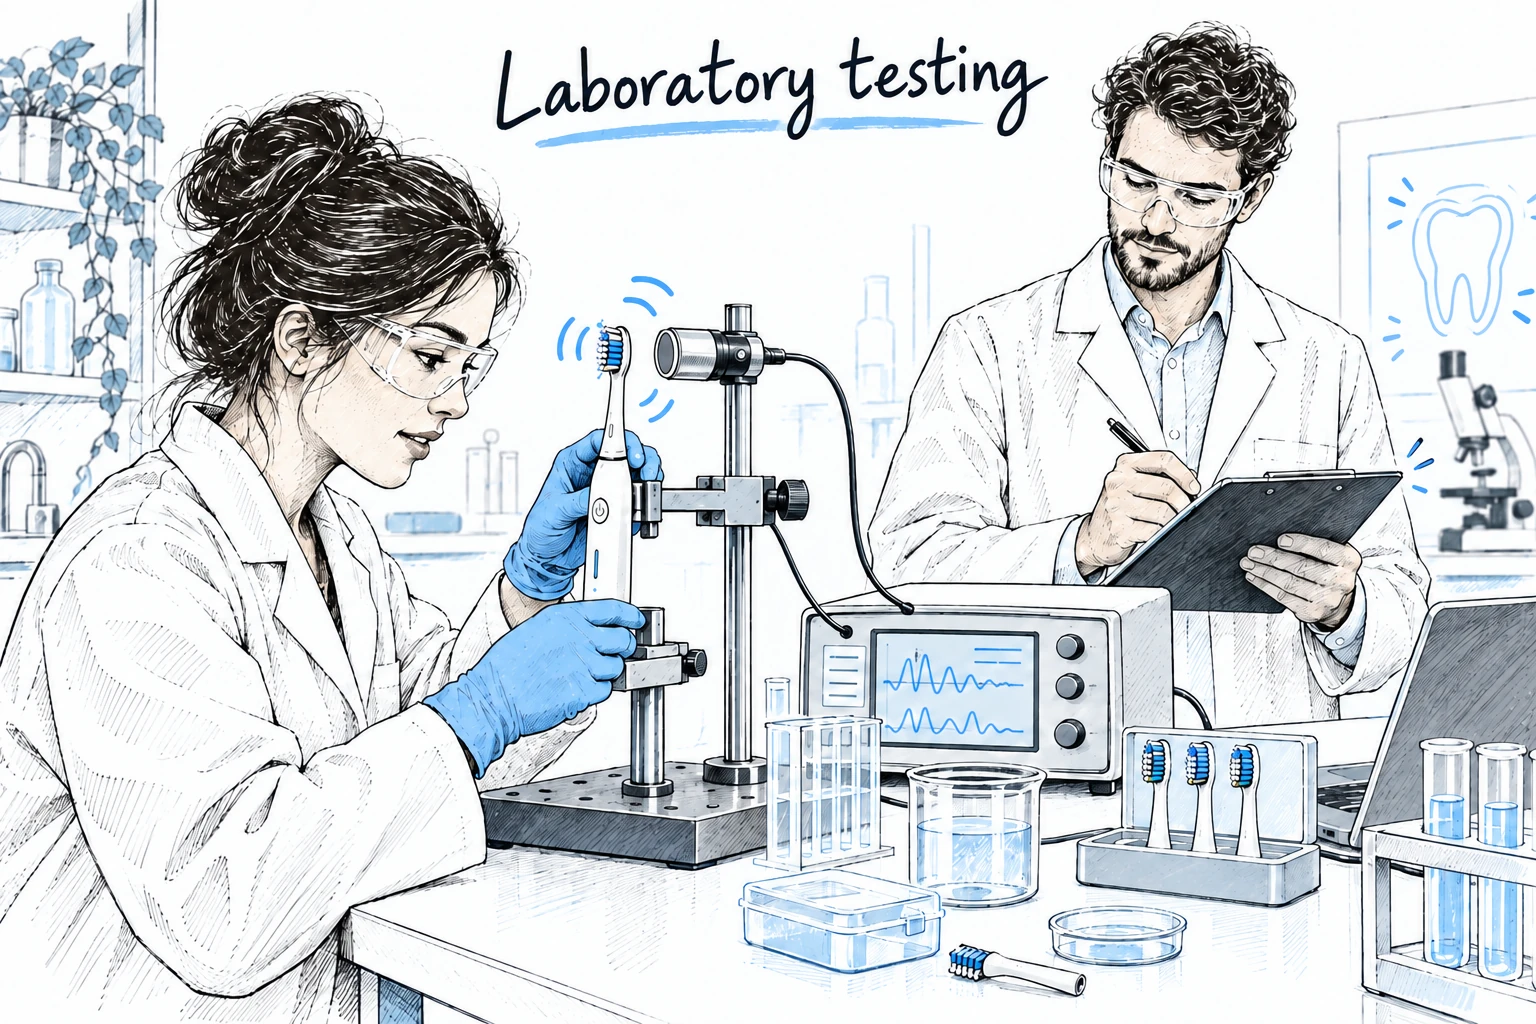
Conduct Laboratory Testing
Build a test plan around the final configuration. List electrical-safety and EMC tests for the requirements that apply, and collect RoHS evidence for restricted substances at homogeneous-material level. Identify models, accessories, test methods, samples, supplier declarations and any targeted laboratory tests.
Clinical Evaluation (MDR Devices Only)
For a product within the MDR, prepare a clinical evaluation using appropriate clinical data and a reasoned assessment of safety and performance. ISO 14155 addresses clinical investigations involving human subjects; it is not the standard for every clinical evaluation. A clinical investigation is needed only when required to address the evidence gap and the applicable rules.
Maintain Quality Processes
Maintain quality processes appropriate to the manufacturer's legal obligations and product. A particular ISO certificate is not automatically required for every consumer product or every MDR class; where a notified-body route or other rule requires a quality system, confirm the specific scope and evidence.
Notified Body Review (if required)
Use a notified body when the applicable conformity route requires one. The body assesses the relevant scope under its designation; any certificate is limited to that scope and does not replace the manufacturer's other obligations.
Issue Declaration of Conformity
For equipment covered by the EMC Directive or RoHS Directive, the manufacturer must retain the EU Declaration of Conformity and technical documentation for 10 years after the equipment is placed on the market. Check which legislation applies to each product and which economic operator holds each record duty; other rules may set different periods.
Affix CE Mark
Affix the CE mark only when required and after the applicable conformity assessment is complete. Follow the placement and size rules for the specific legislation, including any exceptions; add a notified-body number only when the route requires it.
Establish Post-Market Surveillance
Maintain a post-market process that records customer complaints, affected batches, nonconforming product and corrective actions. Define how the team investigates issues, traces affected lots, documents the outcome and escalates safety incidents for reporting under the applicable rules.
Manufacturer-Led Assessment and Notified Body Routes
Some conformity routes require a notified body, including certain MDR classifications. Identify the product classification and route before comparing assessment needs; the table below outlines the main routes relevant to this product.
| Factor | Manufacturer-Led Route | Notified Body Route |
|---|---|---|
| Route requirement | Manufacturer-led assessment where the applicable legislation permits it and no notified-body route is required | Required only for conformity routes that specify a notified body, including many higher-risk MDR devices and certain Class I categories |
| Timing | Set the schedule after confirming sample readiness, test plan and any external review. | Confirm the application, review and audit schedule with the notified body for the selected route. |
| Cost | Project-specific; confirm test and documentation costs for the product | Project-specific; obtain a current quote for the required assessment |
| What it demonstrates | The manufacturer documents conformity and remains responsible for the product | Independent assessment within the notified body's designated scope, when required |
| Quality-system assessment | Maintain the quality processes and records required by the applicable legislation. | Notified-body quality-system assessment and audits where required by the selected route. |
| Product changes | Reassess affected requirements and update technical documentation and the declaration as needed. | Notify the notified body or obtain further assessment when the applicable route requires it. |
For a product that is not subject to a notified-body route, the manufacturer may complete the applicable conformity assessment and issue the required declaration. A retailer's preference for third-party review is a separate commercial condition, not proof that a notified body is legally required.
If the MDR applies, use the classification rules and conformity-assessment options for the device's actual class and characteristics. Notified-body involvement is mandatory only for routes that require it; do not infer the route from a general medical claim alone.
Clinical Evaluation Requirements Under MDR
A formal MDR clinical evaluation is relevant only when the product is a medical device. The depth of the evaluation should reflect its intended purpose, risks, available evidence and applicable requirements; a consumer toothbrush does not need an MDR clinical evaluation simply because it is an oral-care product.
For an MDR device, the clinical evaluation should assess relevant clinical data and show how the evidence supports the device's safety and performance for its intended purpose. Depending on the case, the evidence may include:
- Literature review: Identify and appraise relevant published data, including its methods, population, device and limitations. Similar-device data must be relevant to the product being assessed.
- Equivalence: If relying on data from another device, establish and document equivalence against the MDR criteria and ensure the manufacturer has the necessary access to the data. Similarity alone does not justify transferring clinical conclusions.
- Clinical investigation: Consider an investigation when existing evidence does not adequately address the relevant questions and the applicable route requires it. ISO 14155 applies to the design, conduct and reporting of clinical investigations involving human subjects.
- Post-market clinical follow-up: Plan PMCF where it is required or justified for the device and its residual uncertainties. The need and scope depend on the device and evidence; it is not a blanket requirement limited to every Class IIa or higher product.
Update the clinical evaluation and supporting records when new evidence, product changes or post-market information affects the assessment. Use qualified regulatory or clinical expertise for an MDR device.
CE and UKCA Marking in Great Britain and Northern Ireland
Great Britain continues to accept CE marking for many product regimes under current recognition arrangements; medical devices have separate transition rules. Northern Ireland has a distinct framework. Check the applicable regime, product type and current guidance before choosing CE, UKCA or CE UKNI marking.
For products placed on the UK market, distinguish Great Britain from Northern Ireland and distinguish general manufactured goods from medical devices. The applicable recognition and transition dates depend on the product and legislation.
- CE and UKCA: For many manufactured goods, CE marking continues to be recognised in Great Britain under current rules. Medical-device transition arrangements have their own conditions and dates.
- Regulatory oversight: The MHRA oversees UK medical-device requirements and registration. UK Approved Bodies and EU Notified Bodies have distinct designations and scopes.
- Documentation: Prepare technical documents and labeling for the target market and applicable UK rules. Check standards against the relevant designated references and product scope.
- Marking: CE, UKCA and CE UKNI have different roles. Follow the placement and labeling rules for the specific product and market.
- Medical-device transition: Certain CE-marked medical devices may remain on the Great Britain market for a defined period under transition provisions. Check device category, conformity route and current conditions.
For a project targeting both EU and UK markets, identify the applicable rules for each destination and agree responsibilities with the brand owner and manufacturer early. Shared evidence may help, but it does not remove separate market obligations.
Planning EU Compliance With an OEM/ODM Partner
Relish Tech works with oral-care brands on OEM/ODM product projects. For a specific launch, agree with the manufacturer which specifications, supplier records, samples and test evidence are available, who will arrange any external assessment, and who is responsible for the required declarations. Learn more about our OEM/ODM projects.
Compliance responsibilities depend on the product and the legal role of each business. Confirm the relevant company records, product documents and available test evidence for the specific project; do not rely on a general supplier statement as proof of conformity.
For an OEM/ODM project, align these areas with the manufacturer and any external assessment providers:
- Project documentation: Align on the product configuration and collect available specifications, drawings and component information for the project file.
- Pre-test planning: Identify open questions about applicable requirements and discuss the proposed test scope with the selected laboratory or qualified adviser.
- Test coordination: If external testing is needed, agree the samples, methods, schedule, costs and report scope directly for the project.
- Component records: Request supplier information for relevant materials and components, then assess whether additional evidence is needed.
- Market labeling: Prepare instructions and labeling for the destinations where the product will be sold, using the language and content rules that apply there.
- Supplier documentation: Confirm which quality, facility and manufacturing records can be shared for the project and what each record covers.
Discuss the product scope, target markets and available project evidence with our team. Requirements and support depend on the specific product and agreed project scope. See our certifications information for the records currently published by Relish Tech.
- MOQ: Standard custom orders start at 1,000 units; smaller quantities may be considered case by case.
- Schedule: Testing, documentation and production schedules depend on the product, evidence and project scope. Confirm current timing before placing an order.
- Quality controls: Agree inspection criteria, sampling and AQL in the project specification or contract.
- Compliance documents: Confirm which product-specific records and external test reports are available for the project. No single certification or report establishes conformity with every applicable rule.
Common Pitfalls in EU Market Entry for Oral Care Devices
Plan EU market entry by assigning each requirement to the responsible business and aligning product design, documentation and evidence before launch.
- Planning documentation and testing: Prepare the technical information and test samples for the selected assessment route. Coordinate document review and laboratory work with the organizations involved, and make sure each receives the records needed for its part of the process.
- Standards editions: Check the applicable harmonized standards and their current EU references for the product and legislation. A new international edition is not automatically harmonized, and an older edition does not by itself prove non-compliance; assess the applicable legal route and evidence.
- Battery evidence: UN 38.3 concerns battery transport testing and is separate from whole-product CE conformity assessment. Identify the battery rules and transport records that apply to the product and shipment. Explore our electric toothbrush range for product information.
- EMC planning: Where EMC requirements apply, an engineering review or pre-compliance test may help identify issues before formal assessment. The need and scope depend on the final configuration and test plan.
- RoHS evidence: Collect current evidence from suppliers and assess the product at the material level, including applicable limits and exemptions. Use additional testing where the evidence or risk assessment indicates it is needed.
- Language requirements: Provide instructions and safety information in the languages required for the destination markets and applicable product rules. Requirements vary by product and country; they do not mean every SKU needs all 24 EU official languages.
- After-market duties: Maintain the post-market processes required by the applicable product legislation. Define how feedback, complaints and safety issues are recorded and escalated, and distinguish consumer-product duties from MDR vigilance.
Key Takeaways
- Check which EU product legislation applies before determining whether CE marking is required.
- Identify the rules for the product's intended purpose and configuration. EMC, voltage, RoHS, GPSR and MDR have different scopes.
- A toothbrush is not automatically an MDR Class I device. Classification and notified-body involvement depend on the applicable legal route.
- Discuss project scope, evidence owners, testing dependencies and the working schedule early, while design decisions can still be adjusted.
- Great Britain, Northern Ireland and the EU have distinct marking and transition rules; check the destination and product regime.
- Agree document, testing and labeling responsibilities with the brand owner, manufacturer and any external assessment provider.
Discuss EU Product Requirements for Your Toothbrush
Not sure which rules apply to your product? Share its intended use, configuration and target markets with the project team to identify the questions that need a product-specific assessment.
Request a Compliance AssessmentPlanning an EU Market Launch?
If you are developing an oral-care product for the EU, we can discuss the OEM/ODM project scope, available product information and the records needed for your next assessment step.
Contact Our TeamFrequently Asked Questions
References
- European Commission. "Regulation (EU) 2017/745 on medical devices." Official Journal of the European Union. https://eur-lex.europa.eu/eli/reg/2017/745/oj
- European Commission. "Directive 2014/30/EU on electromagnetic compatibility." Official Journal of the European Union. https://eur-lex.europa.eu/eli/dir/2014/30/oj
- European Commission. "Directive 2014/35/EU on electrical equipment designed for use within certain voltage limits." Official Journal of the European Union. https://eur-lex.europa.eu/eli/dir/2014/35/oj
- European Parliament and Council. "Directive 2011/65/EU (RoHS) on the restriction of the use of certain hazardous substances in electrical and electronic equipment." Official Journal of the European Union. https://eur-lex.europa.eu/eli/dir/2011/65/oj
- British Standards Institution. "BS EN ISO 14971:2019 Medical devices — Application of risk management to medical devices." BSI Standards Limited.
- European Commission. “MDCG 2019-15 rev.1: Guidance notes for manufacturers of Class I medical devices.” Official guidance and document: MDCG 2019-15 rev.1
- UK Government. “Regulating medical devices in the UK.” Regulating medical devices in the UK
- European Commission. “CE marking: placing products on the EU market.” EU CE marking guidance
- European Parliament and Council. “Regulation (EU) 2023/988 on general product safety.” General Product Safety Regulation
- European Commission. “EUDAMED: first four modules mandatory from 28 May 2026.” European Commission update
- U.S. Food and Drug Administration. “Product Classification: Powered toothbrush (JEQ, 21 CFR 872.6865).” FDA Product Classification